With advances in structural biology and high-performance computing, molecular dynamics simulations play an increasingly important role in drug discovery. These simulations can provide detailed insights into the dynamics and interactions of drug molecules and their biological targets. This information can inform optimization of ligand series and accelerate the development of new therapeutics.
Molecular dynamics can advance your project by:
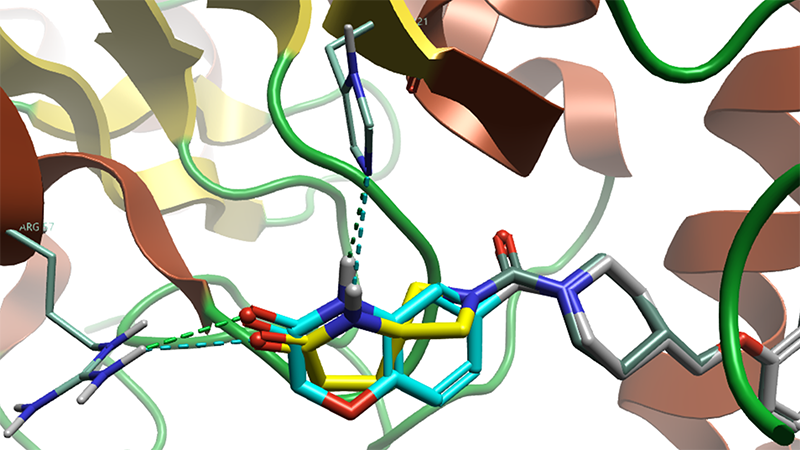
- Understanding drug-target interactions: unlike static structural snapshots, a simulated trajectory provides a statistical description of the key ligand-protein interactions, allowing for a more representative evaluation of binding.
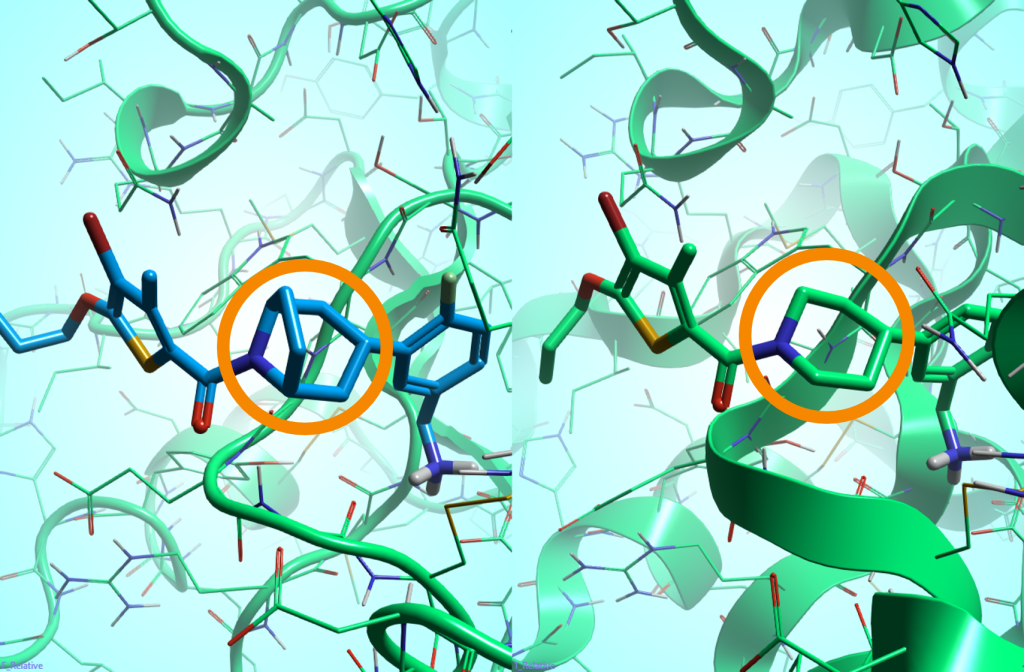
- Identifying conformational changes: a simulation can capture subtle conformational rearrangements of the protein upon ligand binding, or upon modifying the ligand in the active site, which can inform rational design of new ligands.
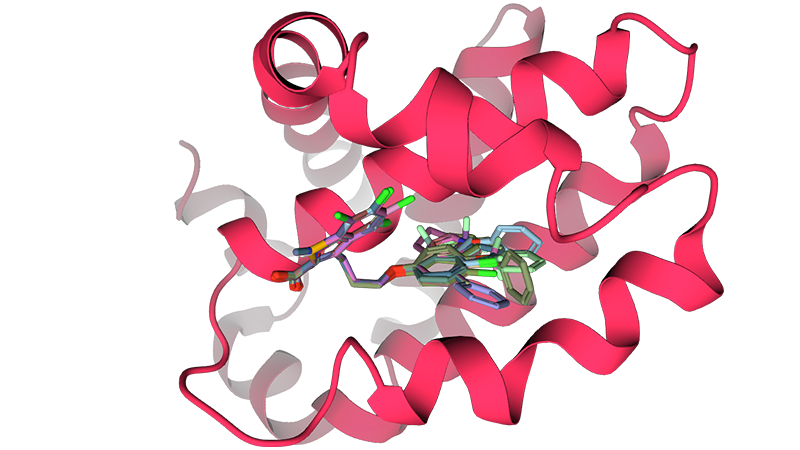
- Validating docking results: by simulating the docked complexes, one can assess the stability of diverse ligand poses, thereby identifying the most likely binding mode(s).
- Enhancing binding free-energy predictions: molecular dynamics can be applied to generate models or ensembles that serve as high-quality inputs for binding free-energy calculations, such as those by Free Energy Perturbation (FEP) or MM/GBSA methods.
- Identifying druggable sites: when combined with Pocket Detection, a molecular dynamics simulation can reveal the statistical accessibility of druggable sites by accounting for protein conformational changes under physiological conditions.
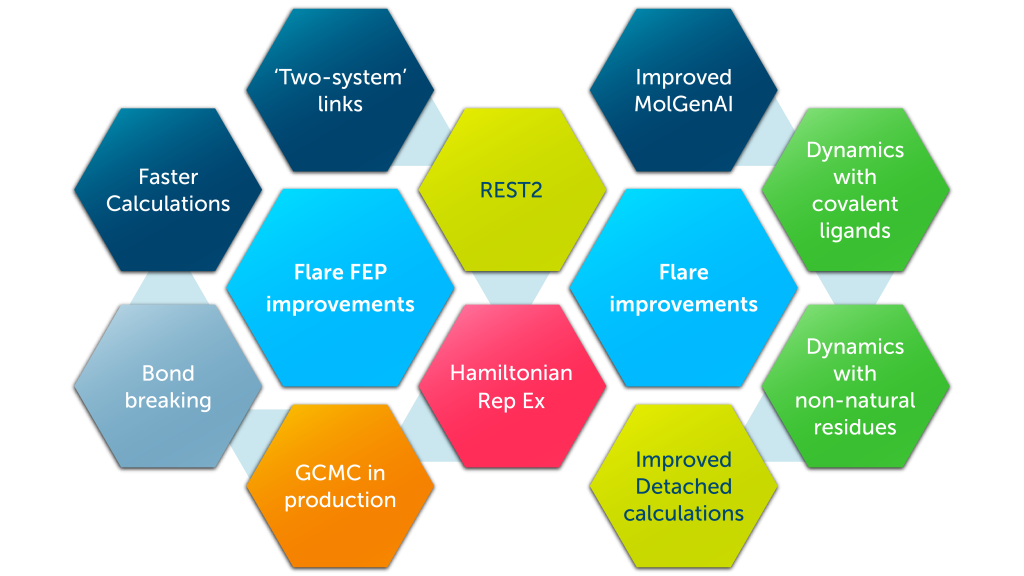
- Accounting for modified residues and covalent ligands: model covalent ligands and proteins with non-natural residues, which can enable accurate characterization of stability and conformational effects in systems that require non-standard force-field parameters.
Molecular dynamics simulations provide a powerful approach to predict and optimize drug-target interactions. The application of molecular dynamics for analysis and design allows the dynamic nature of a system to be better represented, compared to use of a static representation of the system.